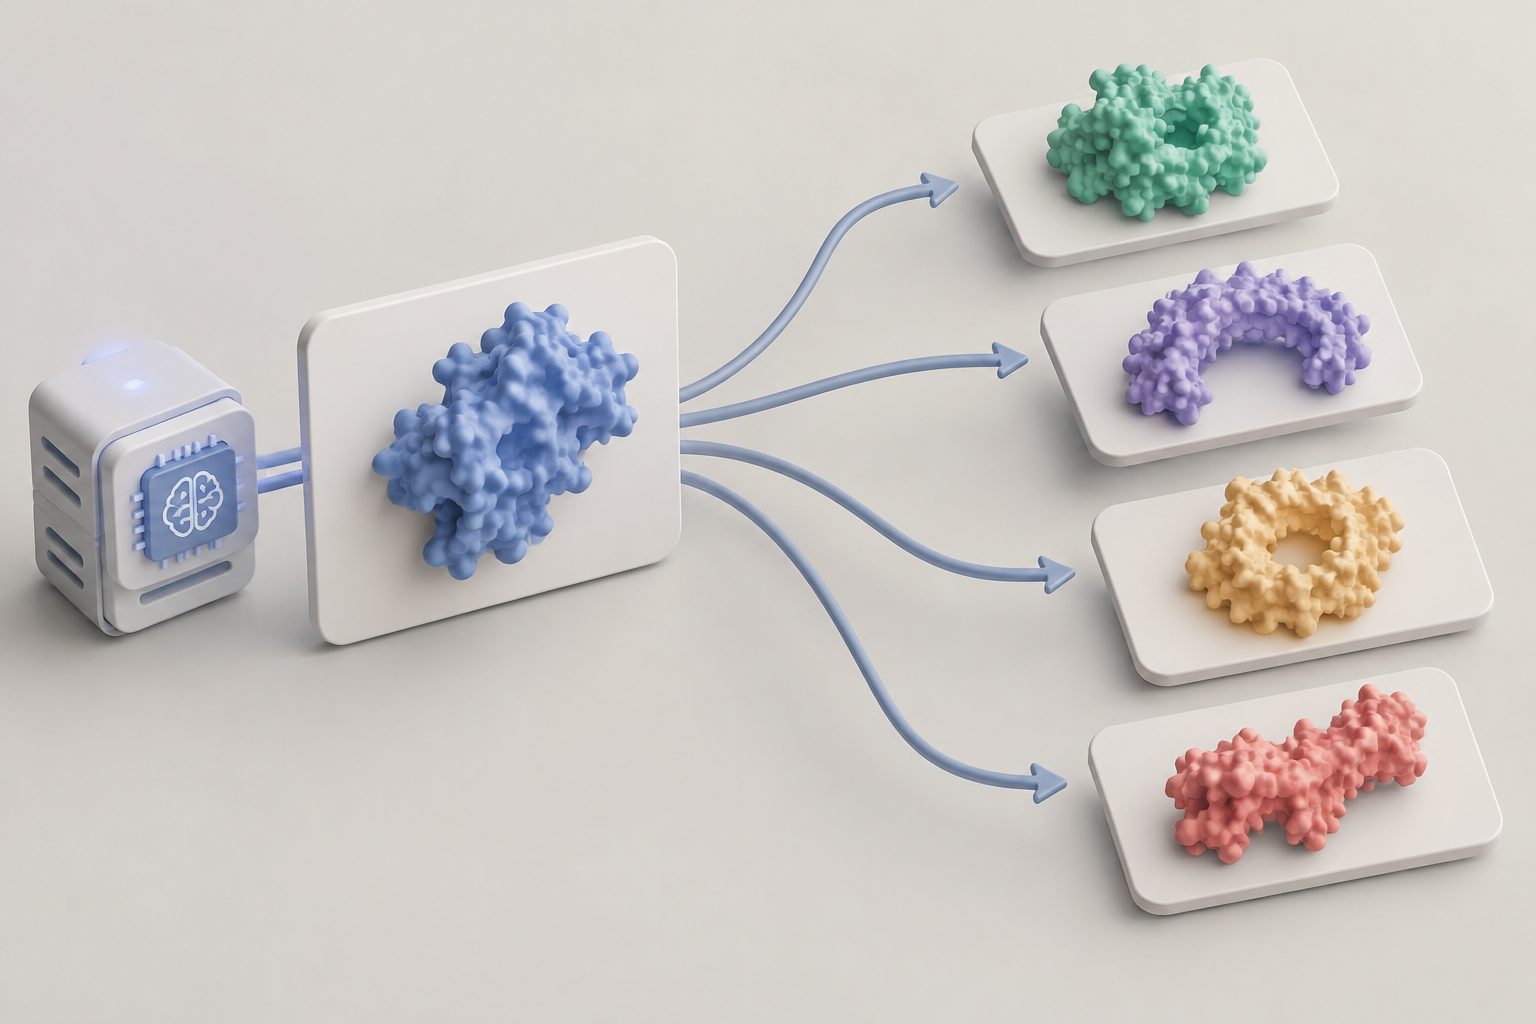
Researchers used distance information inside an AlphaFold2 prediction to model alternative protein conformations.
AlphaFold2 is widely used to predict protein structures, but it often favors one conformation. The researchers found that its predicted distances between pairs of amino acids can contain several likely values, indicating structural heterogeneity before the system completes its refinement process.
They separated these multiple distance patterns into state-specific restraints and added them to the Rosetta modeling system. Across protein systems with different kinds of shape changes, the approach generated alternative conformations from an input structure, with results comparable to those from existing methods.
Finding protein shape clues
The researchers found that AlphaFold2 predictions made with a deep multiple sequence alignment can still encode information about structural heterogeneity. In particular, predicted residue-to-residue distance distributions sometimes showed multiple probability peaks before recycling, the repeated refinement steps used by AlphaFold2.
They fitted and separated these peaks to derive distance restraints for different protein states. When incorporated into Rosetta as energy terms, the restraints allowed the researchers to sample alternative conformations and model large-scale transitions across multiple protein systems. The resulting models were comparable to those produced by existing approaches.
Why extra shapes matter
AlphaFold2 is often used to produce a favored protein structure, but proteins can shift between multiple shapes. This work suggests that information about those alternative states may already be present in a standard prediction and can be used to study protein flexibility and dynamics without relying only on methods that deliberately alter the input sequence information.
The approach could broaden the use of AlphaFold2 for modeling proteins that occupy more than one structural state, including systems that undergo substantial conformational changes.
Evidence and caveats
The evidence comes from computational modeling across multiple protein systems with varied conformational changes. The study compared the generated models with results from existing approaches, and reports that they were comparable.
The abstract does not give the number or identities of the protein systems, the quantitative accuracy of the models, or experimental confirmation that the modeled states occur in nature. The method starts from an input conformation and depends on distance information inferred by AlphaFold2, so its performance may vary across proteins and types of structural change.
// Source
Quantitative Biology · 2026 · DOI: 10.1002/qub2.70062
Authors: Jiaxuan Li, Zefeng Zhu, Chen Song
Institutions: Peking University, Center for Life Sciences, Yantai University