A repository article argues that parts of the genetic code also shape translation speed, protein folding and DNA stability.
The article presents the genetic code as more than a four-letter system for assigning amino acids. It argues that the code’s redundancy also carries information about translation kinetics and co-translational folding, while two chemical property axes—described as S/W and R/Y—encode features related to thermodynamic stability and DNA geometry.
The authors report that 27% of the code’s capacity is degenerate, meaning that different DNA sequences can specify the same amino acid. They calculate that this capacity corresponds to 1.61 bits per codon, or about three effective states, which they say matches the three-part organization of the wobble position. The article also introduces RAD-Diagnostic, a proposed framework for interpreting synonymous variants, with criteria for future testing.
Three proposed DNA layers
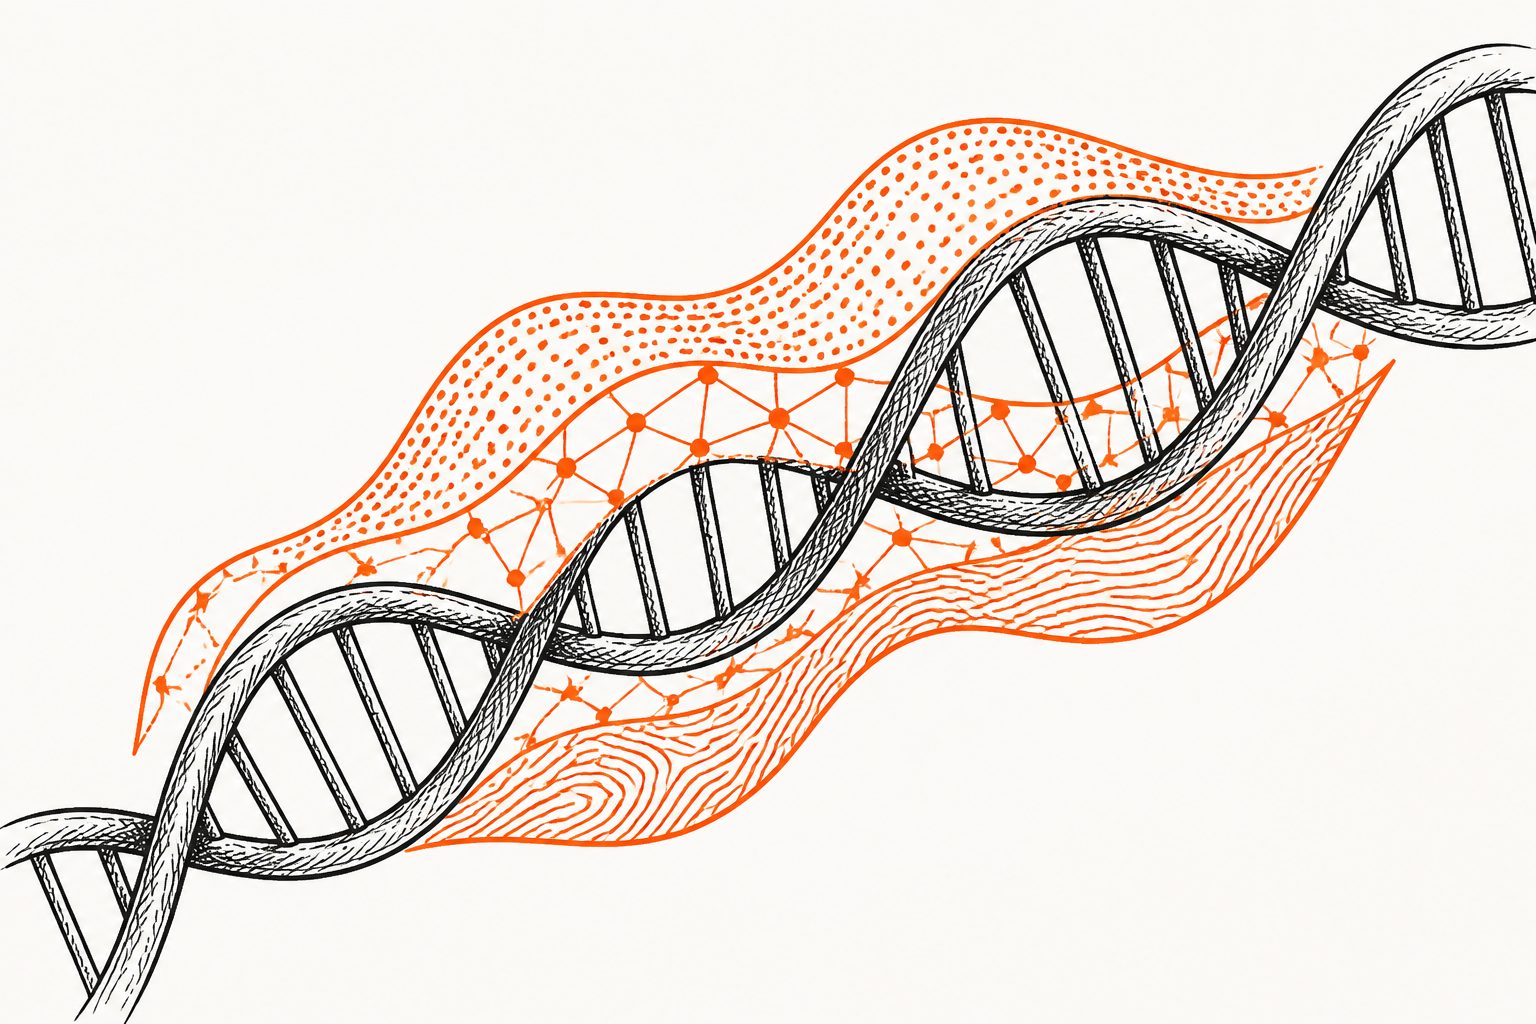
The authors propose three information layers in DNA. The first assigns codons to amino acids. The second is proposed to influence translation speed and protein folding through differences at the wobble position, where some DNA changes do not alter the amino acid. The third is proposed to encode thermodynamic stability and helical geometry through two DNA property axes.
In an analysis of five human genes using composition-matched null models, the authors report that degenerate sites were compositionally controlled in almost all cases examined. BRCA1 was identified as a rare locus-specific exception, with a reported Benjamini–Hochberg-adjusted result of p = 1.3×10⁻⁵. The paper also presents a proposed explanation for how one amino acid sequence can form different stable structures: information in the second and third layers may influence the route by which a protein folds.
Why overlapping information matters
If the proposed framework is supported by future tests, DNA changes that leave an amino acid sequence unchanged could still carry information relevant to translation and protein folding. That could affect how researchers interpret synonymous variants, which are often assessed mainly by whether they change a protein’s amino acid sequence.
The article’s RAD-Diagnostic framework is intended to apply this idea to clinical interpretation of such variants. However, the abstract describes explicit criteria for prospective validation rather than results from that validation, so the clinical usefulness of the framework remains to be established.
Evidence and open tests
The work is an article deposited in the Zenodo repository. Its evidence combines a proposed information model with computational analysis of five human genes and composition-matched null models. The abstract says the calculations can be reproduced from public data using a single script.
The findings are limited by the small number of genes analyzed and by the fact that the abstract does not describe direct experimental tests of translation speed, protein-folding paths, thermodynamic stability or helical geometry. The proposed clinical framework has not yet been prospectively validated in the information provided. The BRCA1 result is reported as an exception, but the abstract does not establish why it occurs.
// Source
Zenodo (CERN European Organization for Nuclear Research) · 2026 · DOI: 10.5281/zenodo.18243694
Authors: Moss Eva